Quality system, risk file and technical documentation
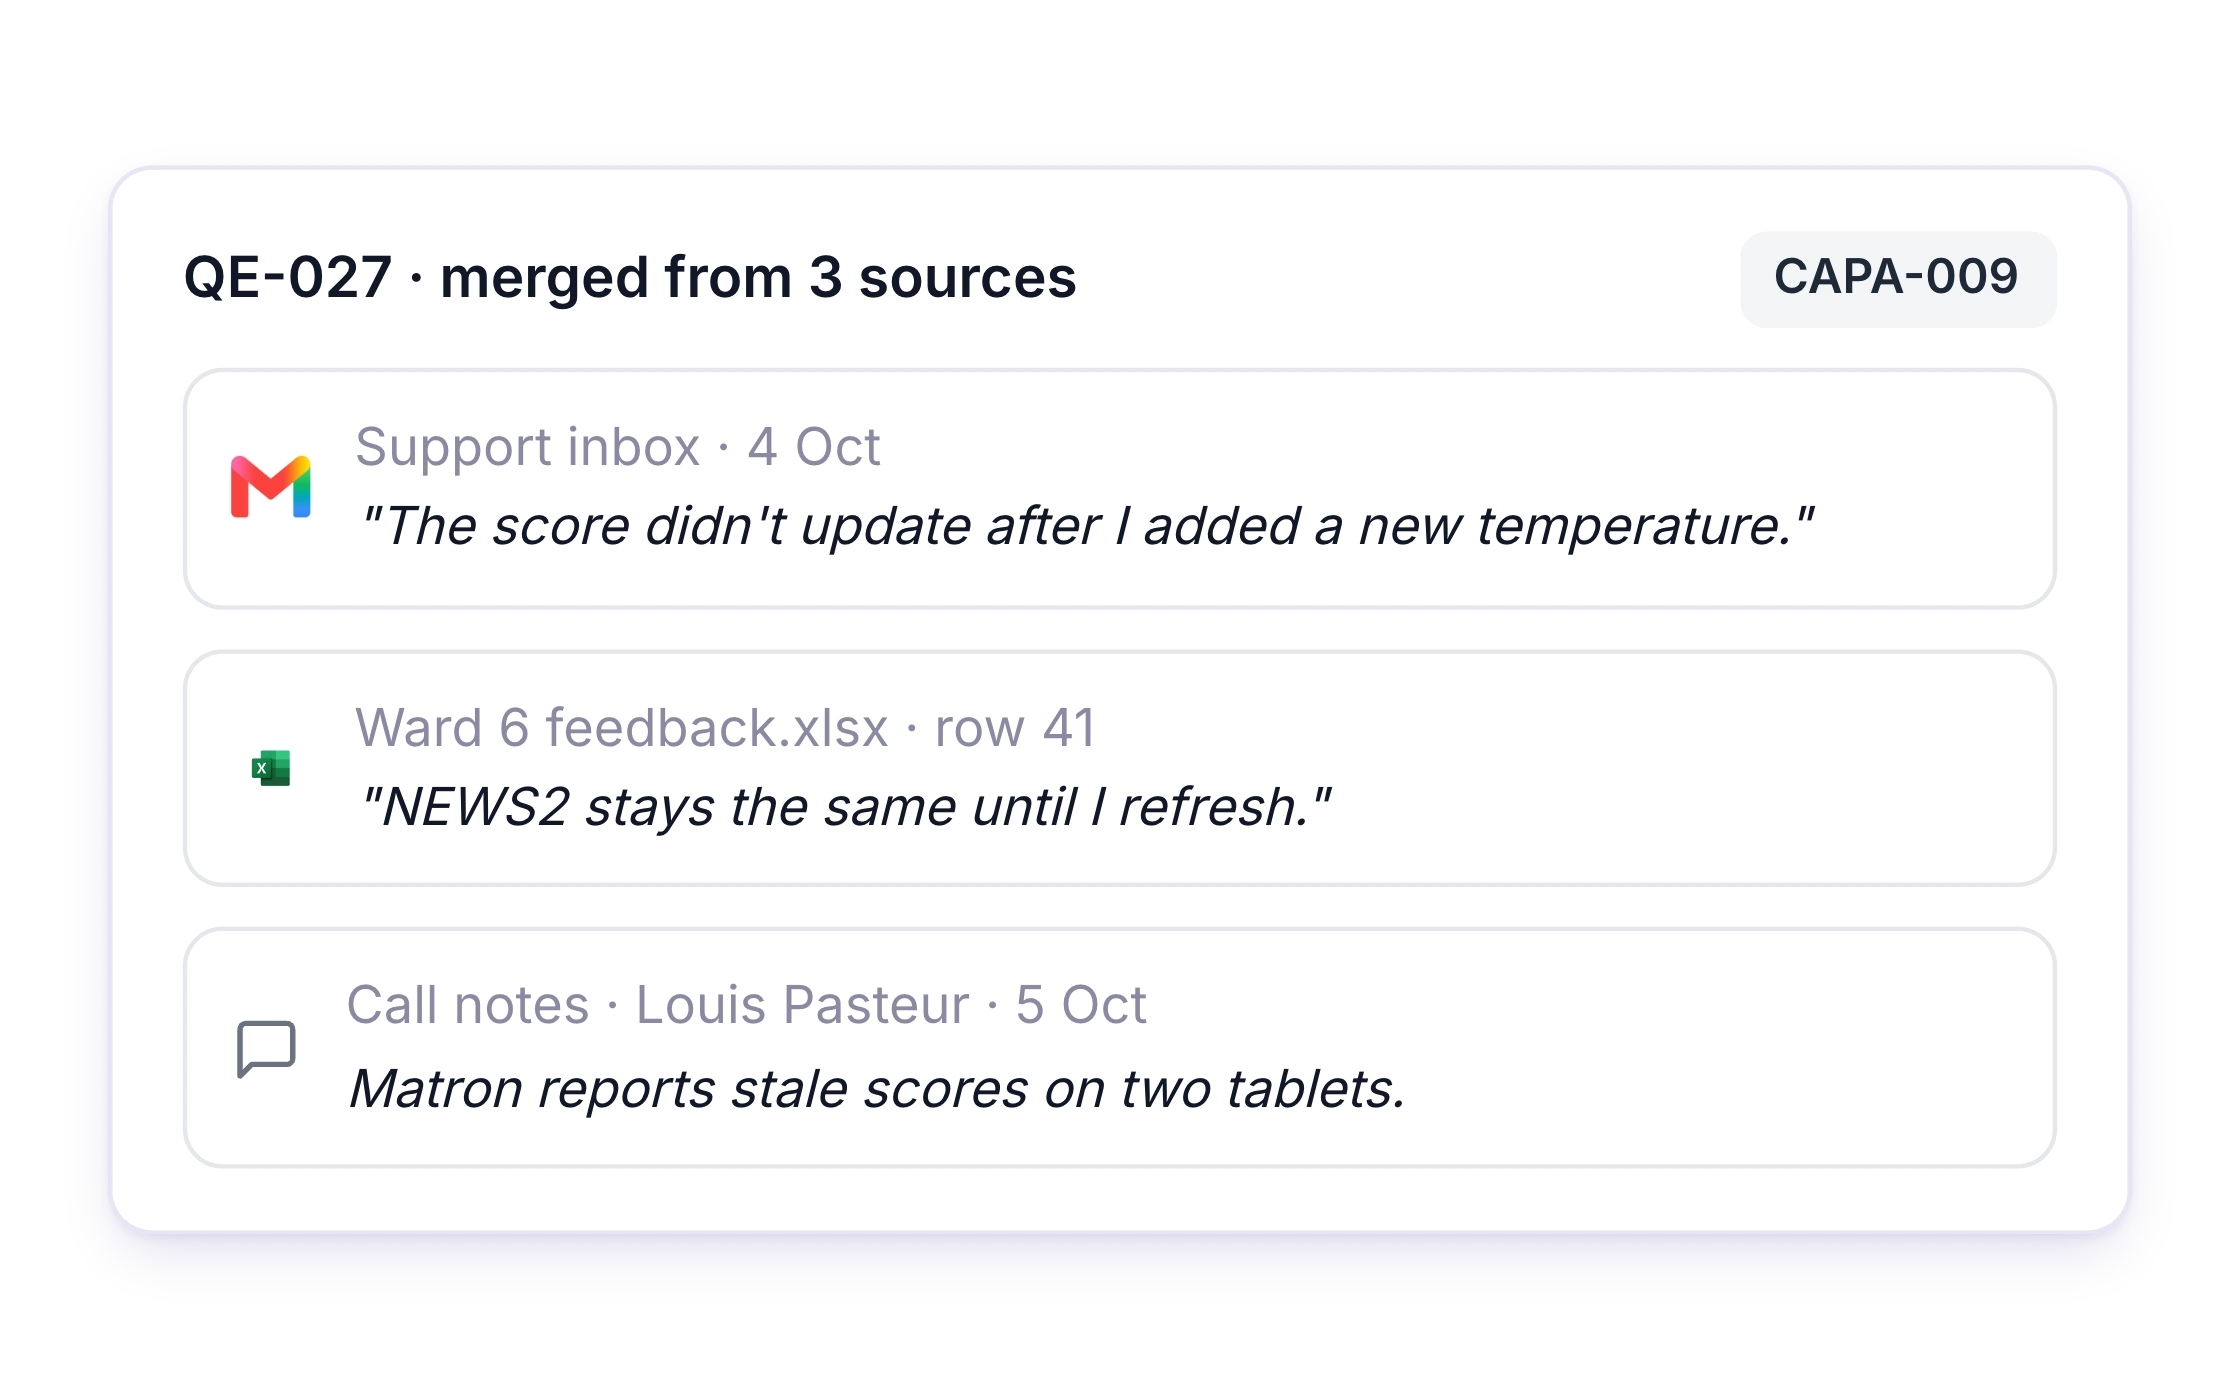
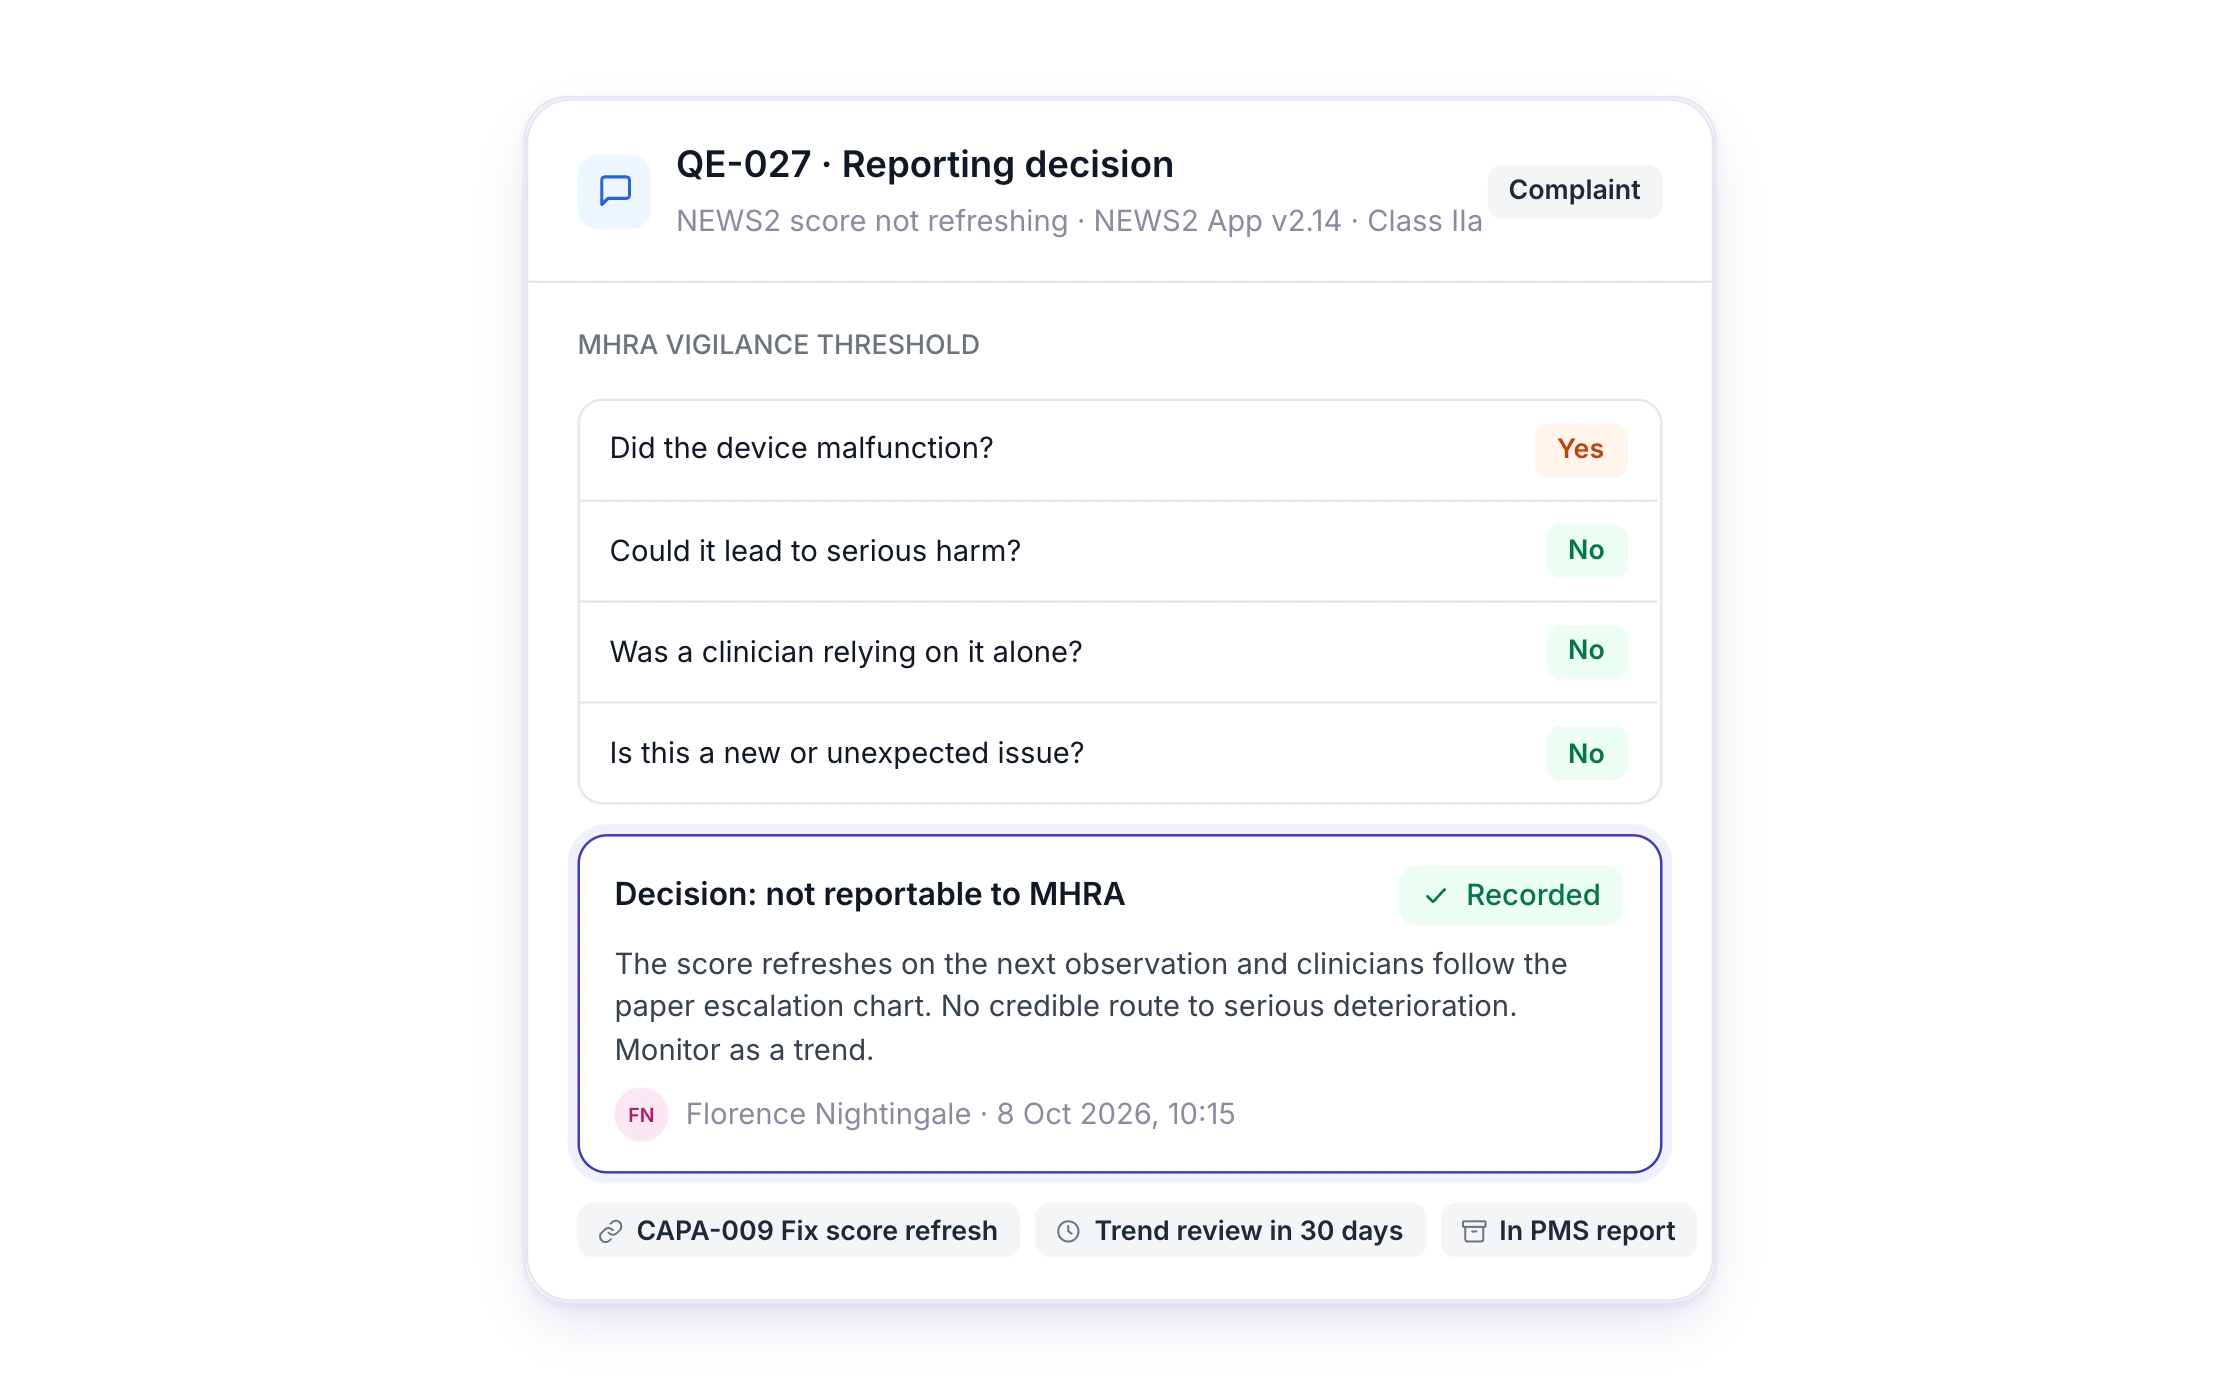
Assuric holds all three key requirements in one place, ensuring a record written once counts for ISO 13485, IEC 62304 and the MDR together rather than being rewritten across systems.
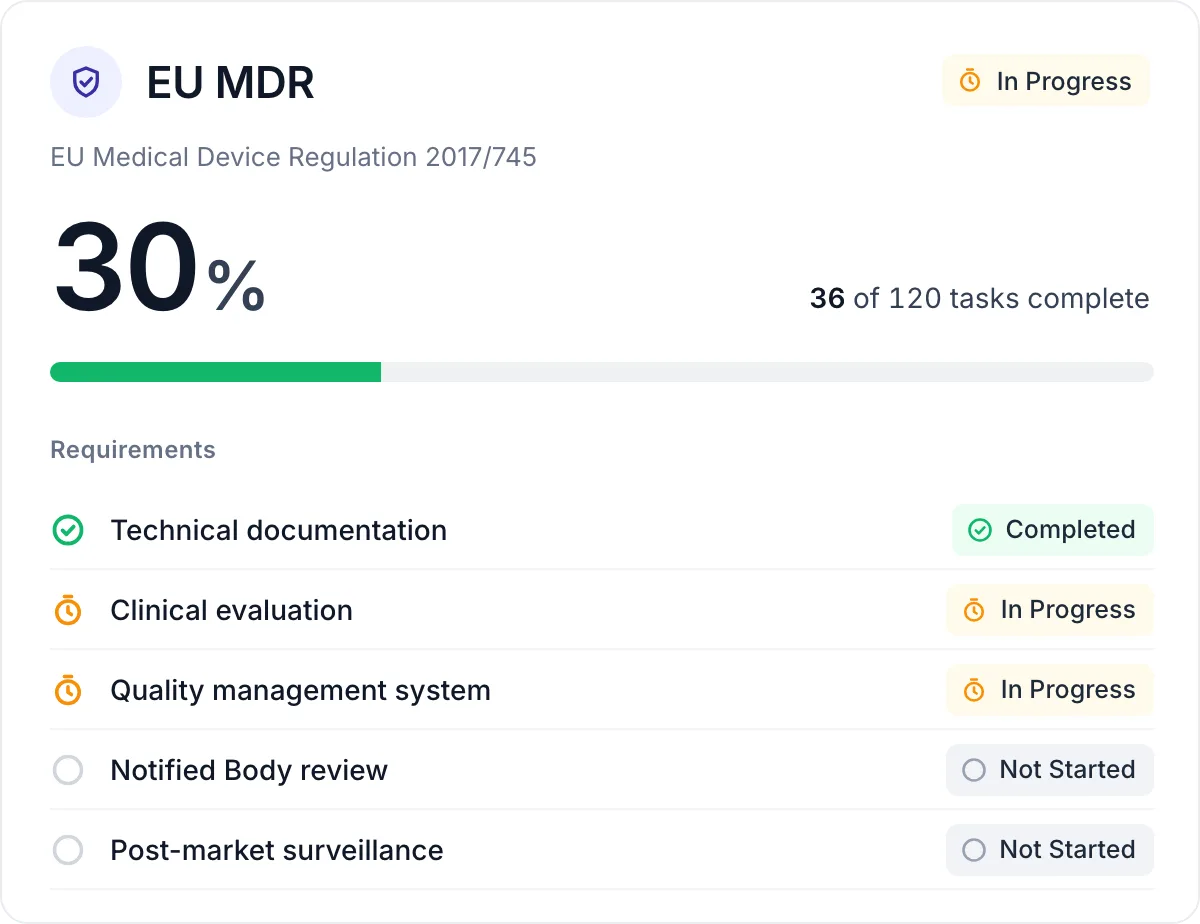
Assuric gives you the QMS, risk file and technical documentation inputs to get through EU MDR conformity assessment, all in one place.
Trusted bymedical device teams


Features
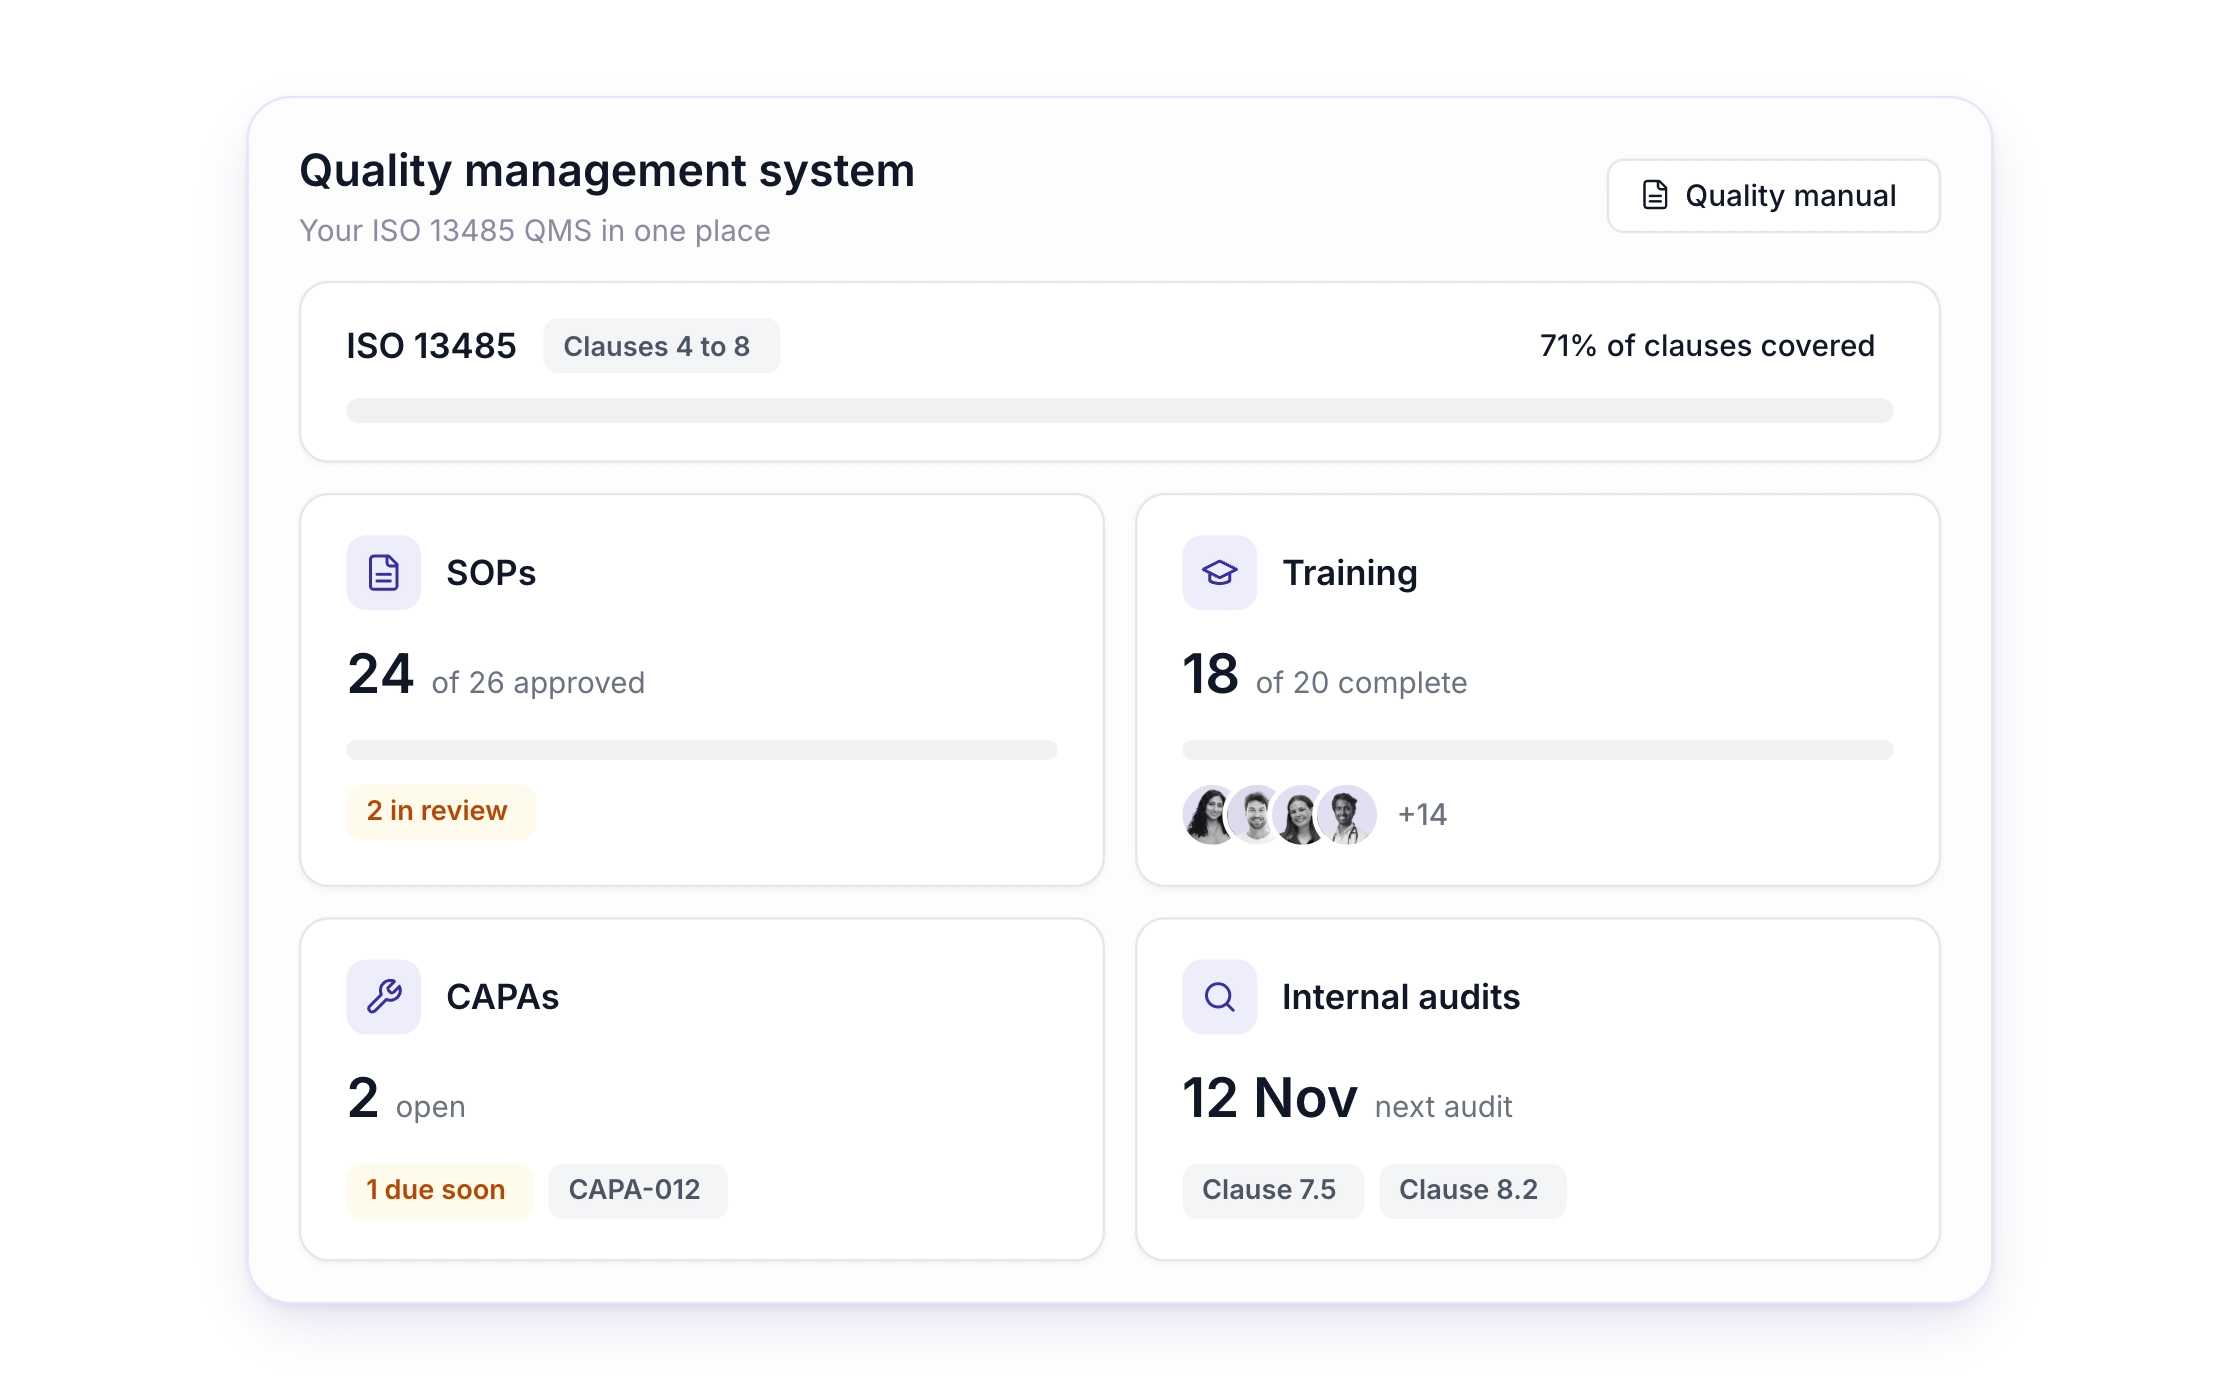
Complete eQMS
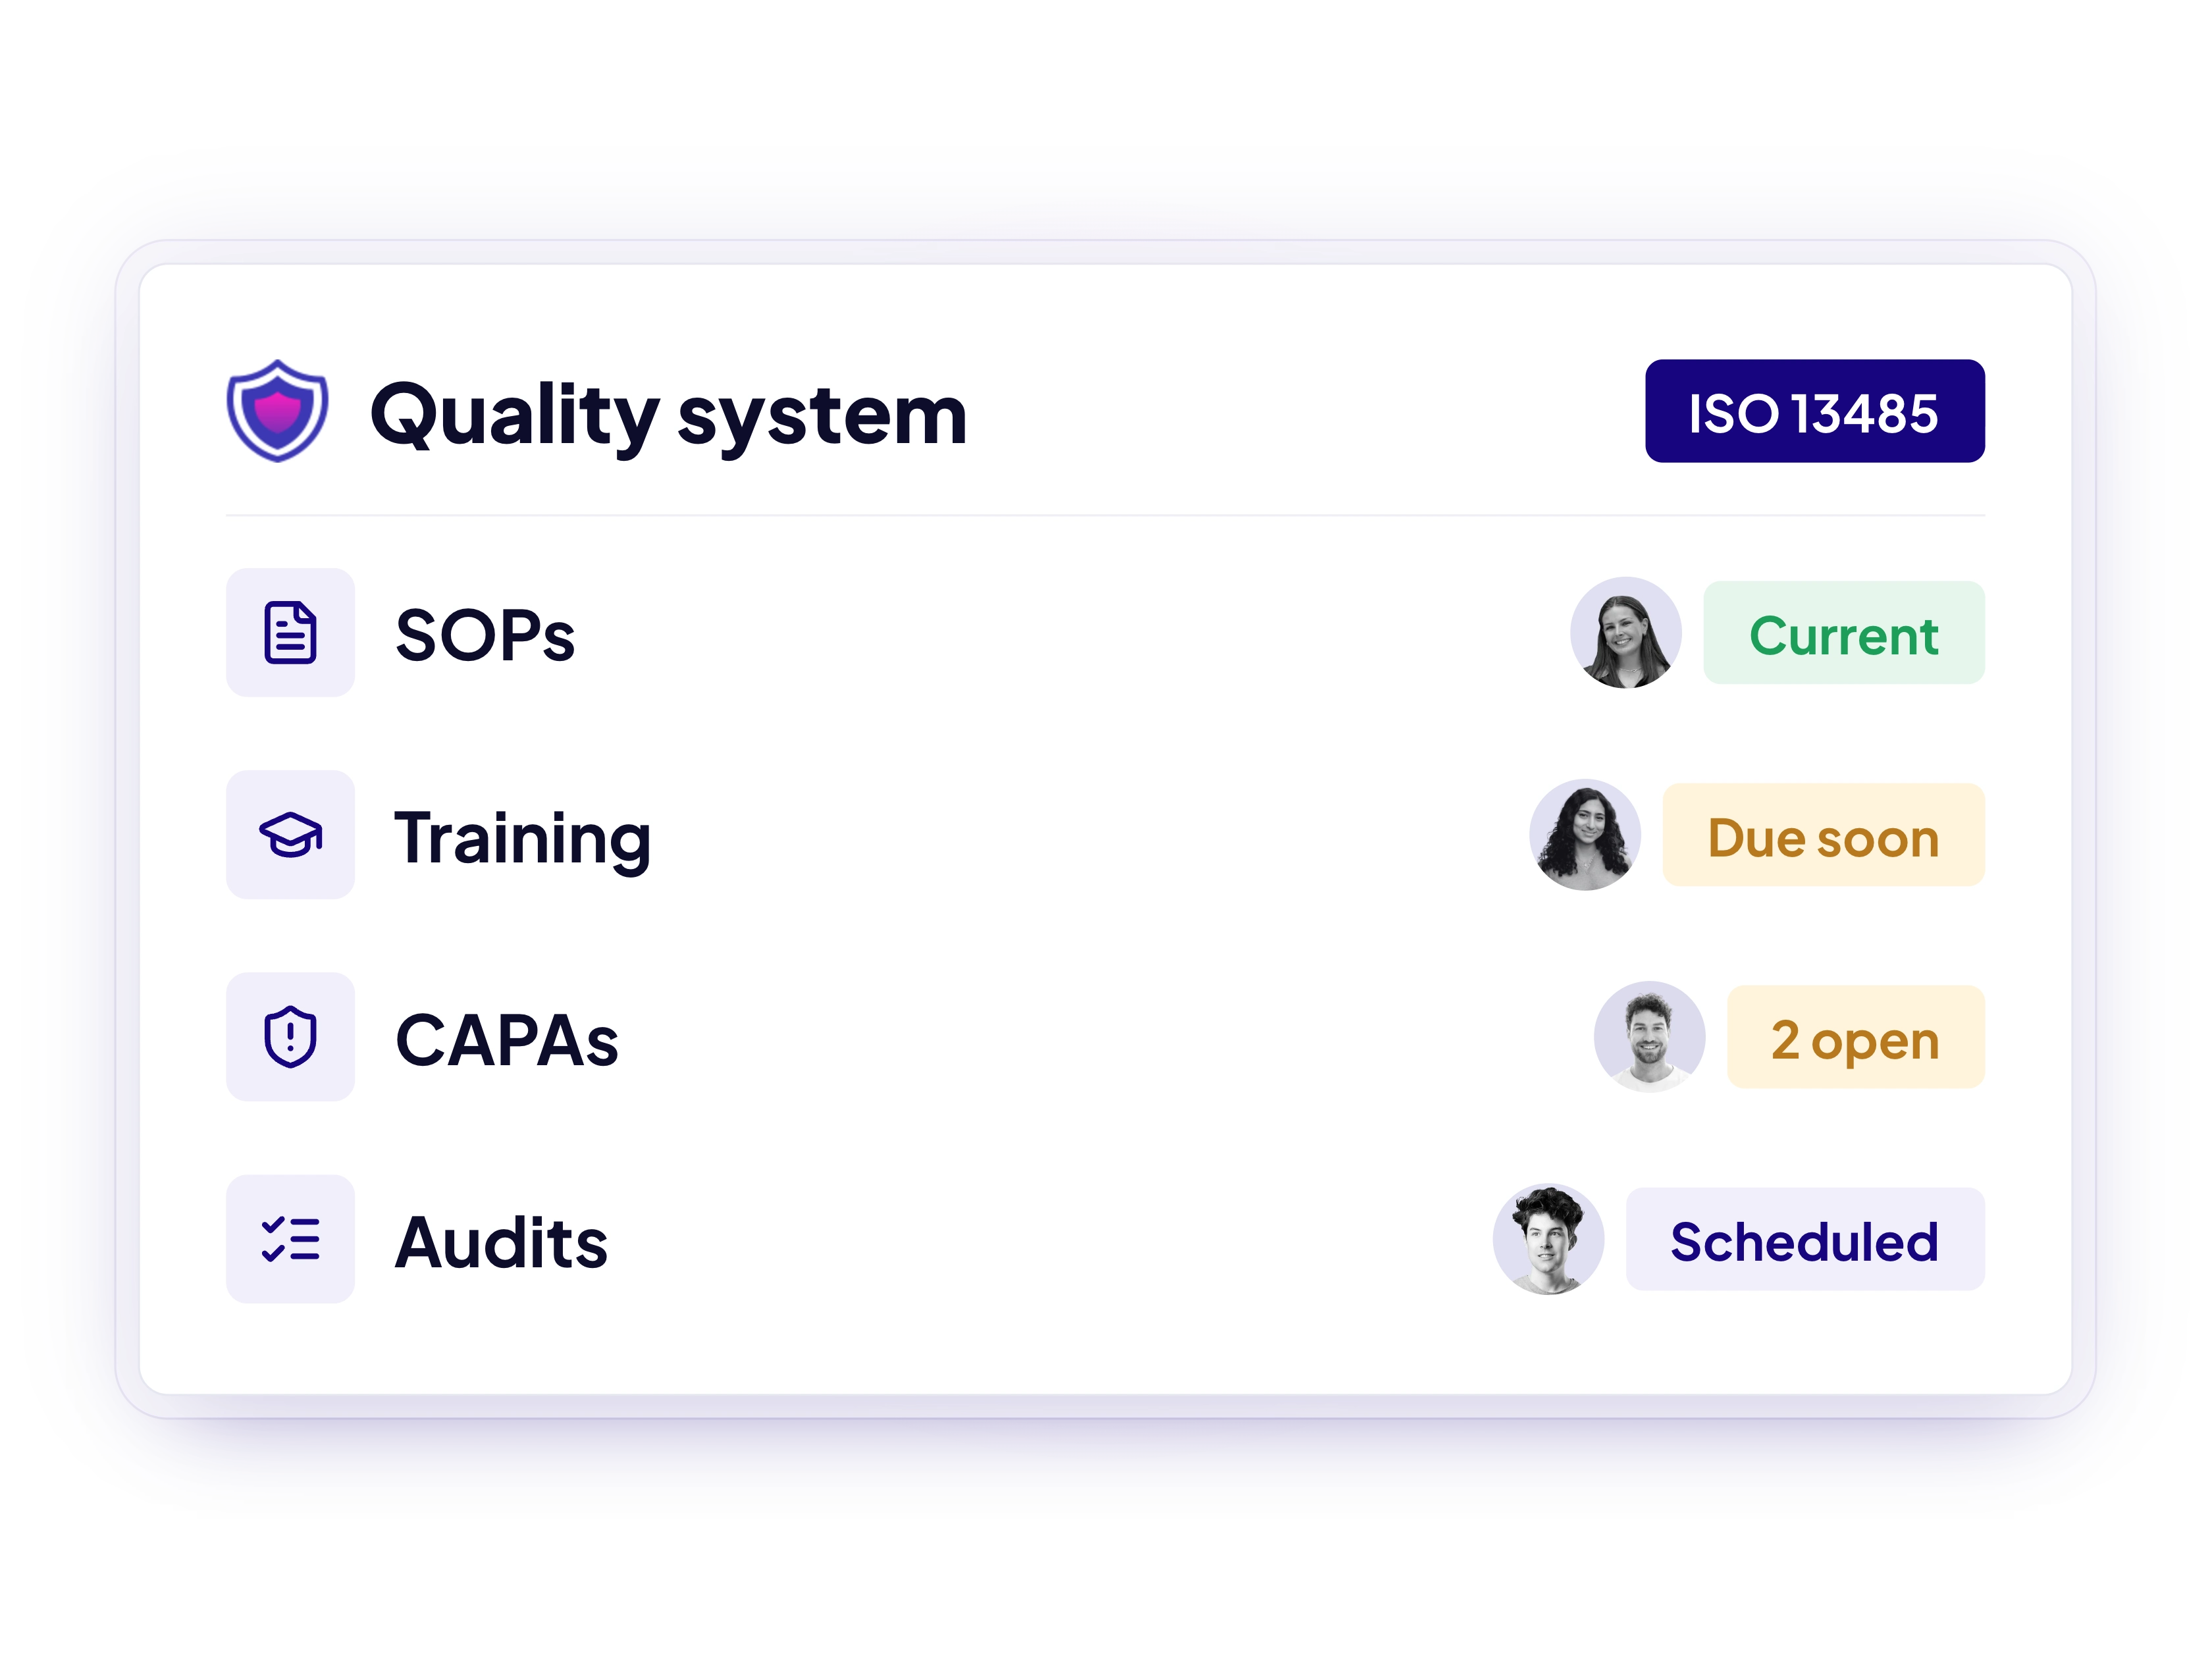
SOPs, training records, CAPAs and internal audits pre-built to ISO 13485, versioned and approved inside the platform, ready to adapt to how you actually work.
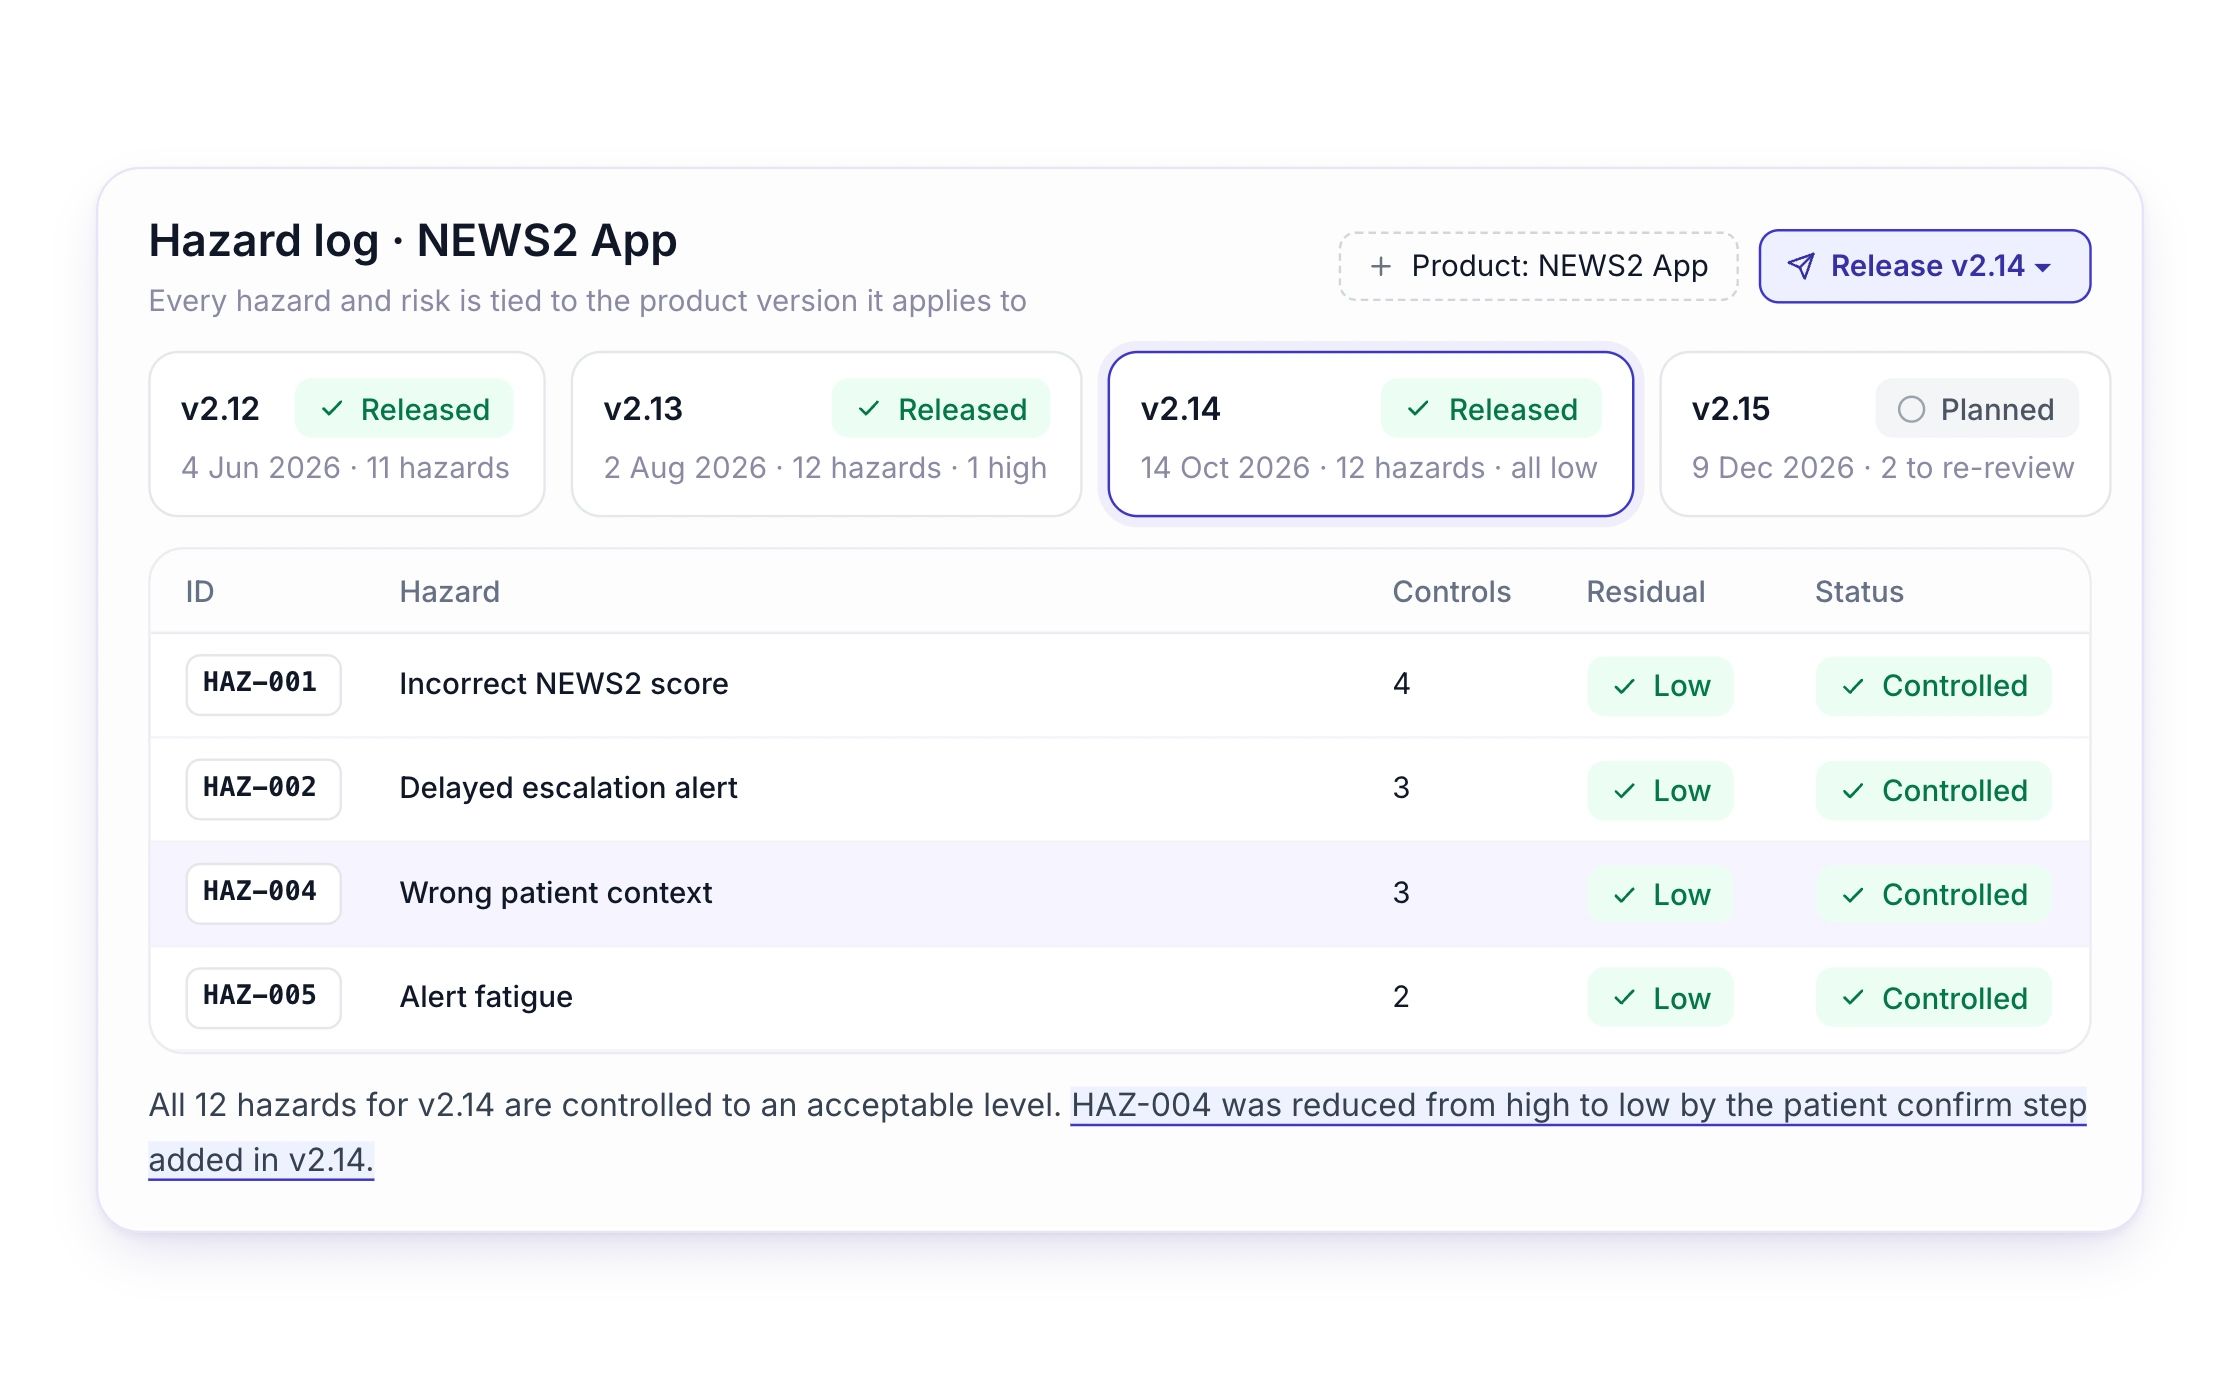
Medical device risk management
Hazards, causes, controls and residual risk in one ISO 14971 risk file, scored and reviewed, and linked to the releases they affect.
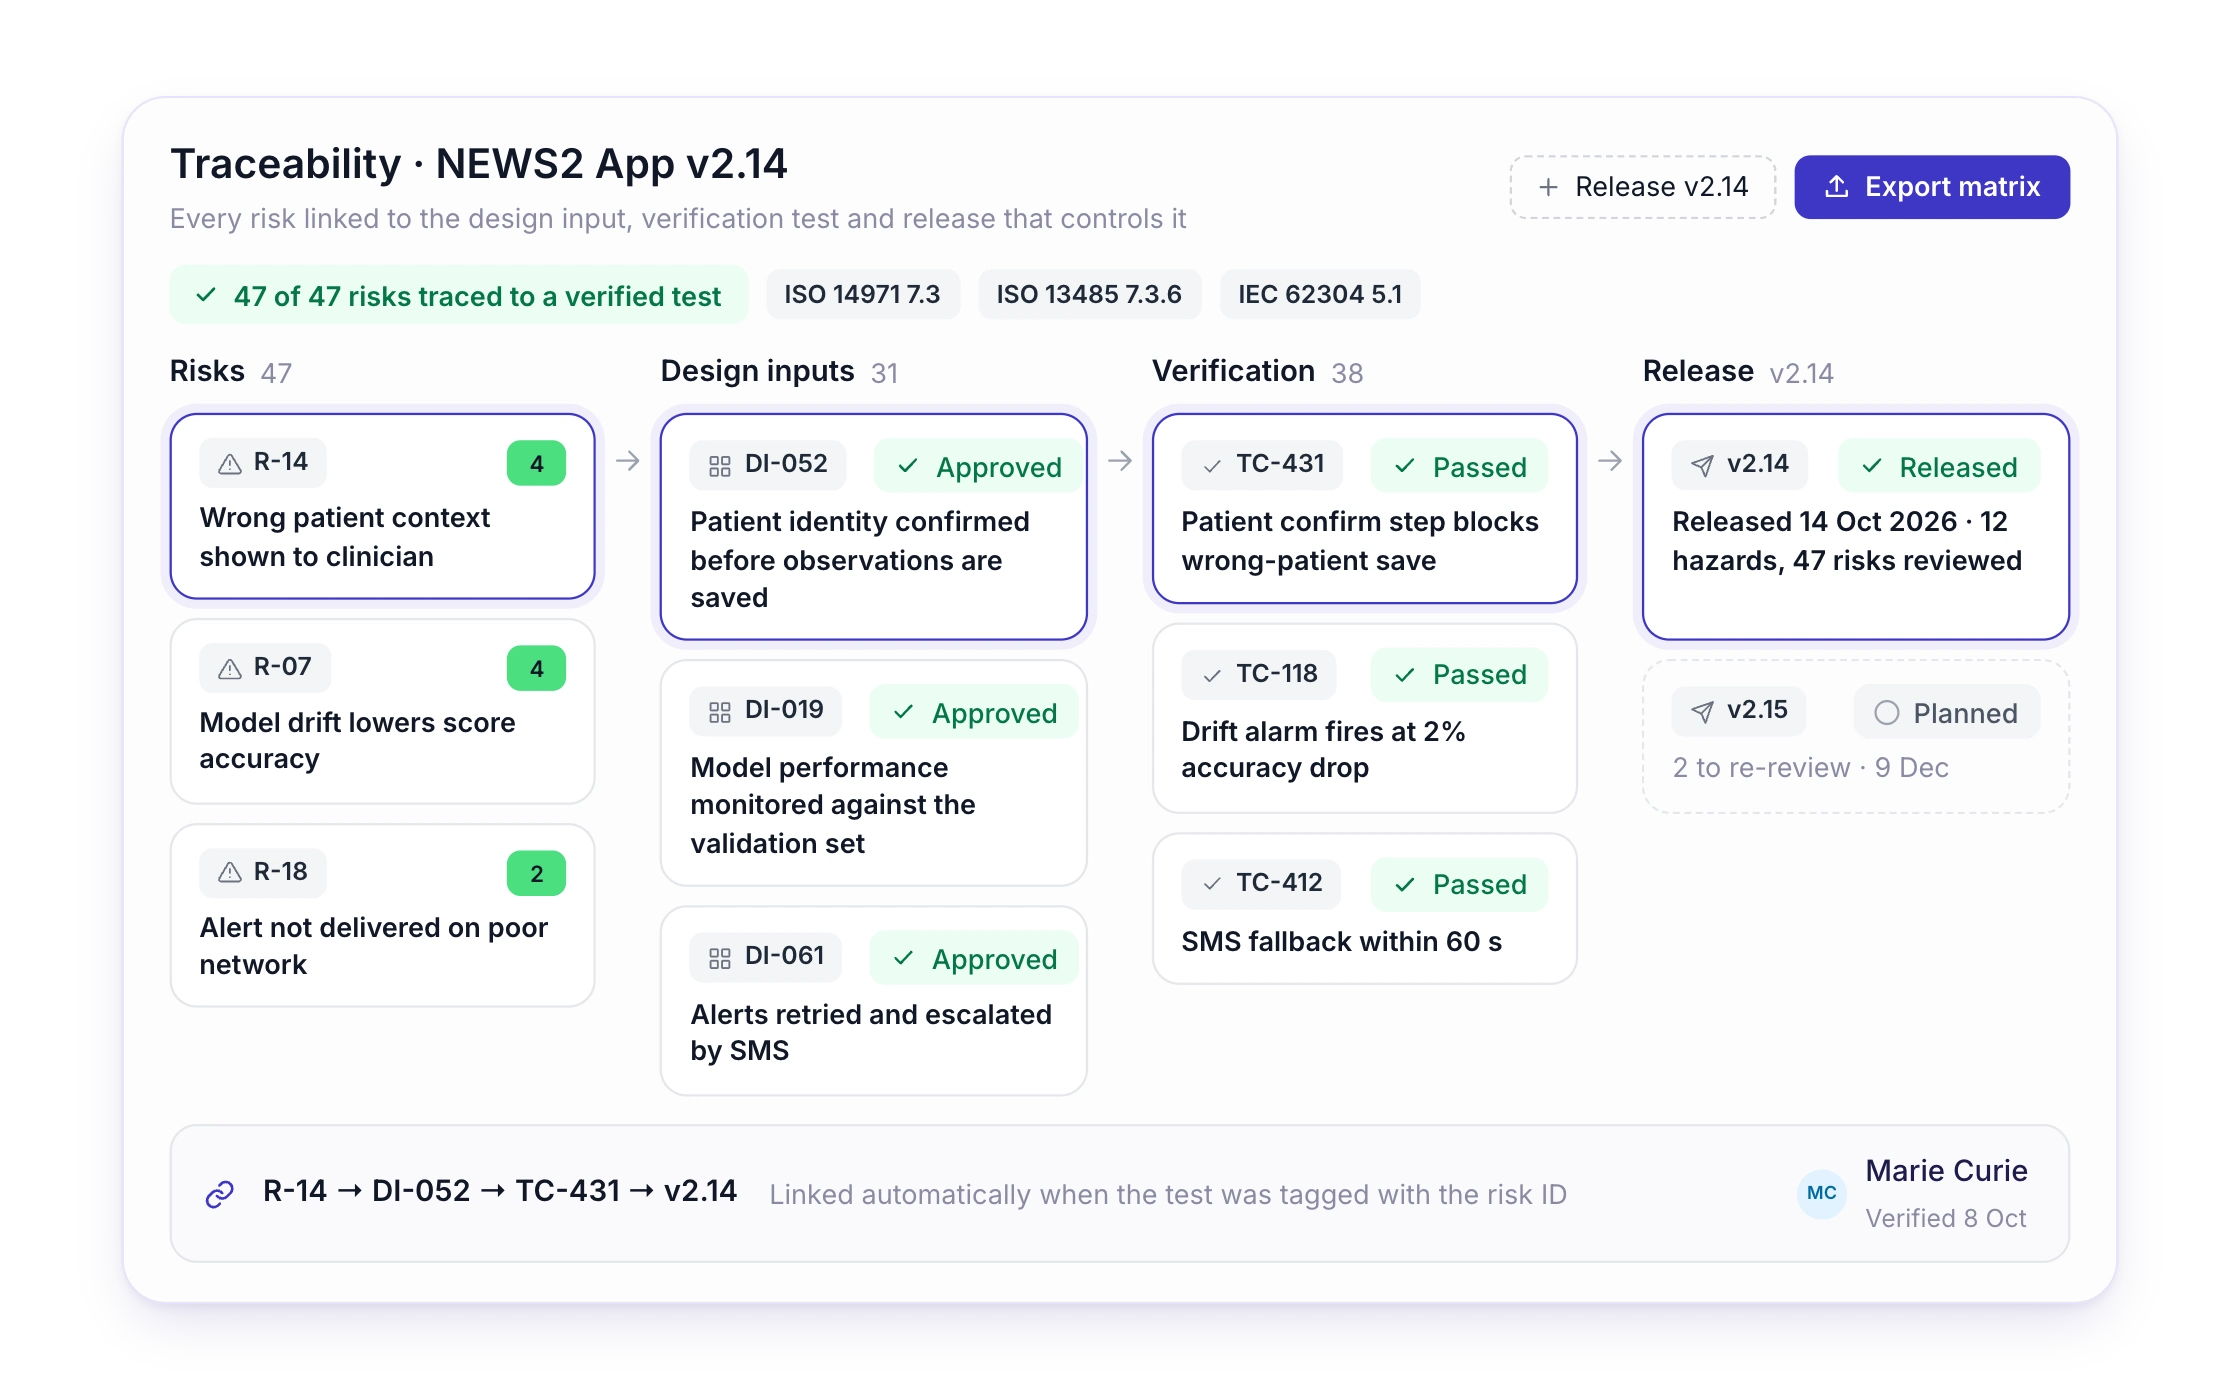
Complete eQMS
Your quality system, risk file and design records all sit in one place, so your technical documentation is built from evidence you already hold rather than started from scratch.
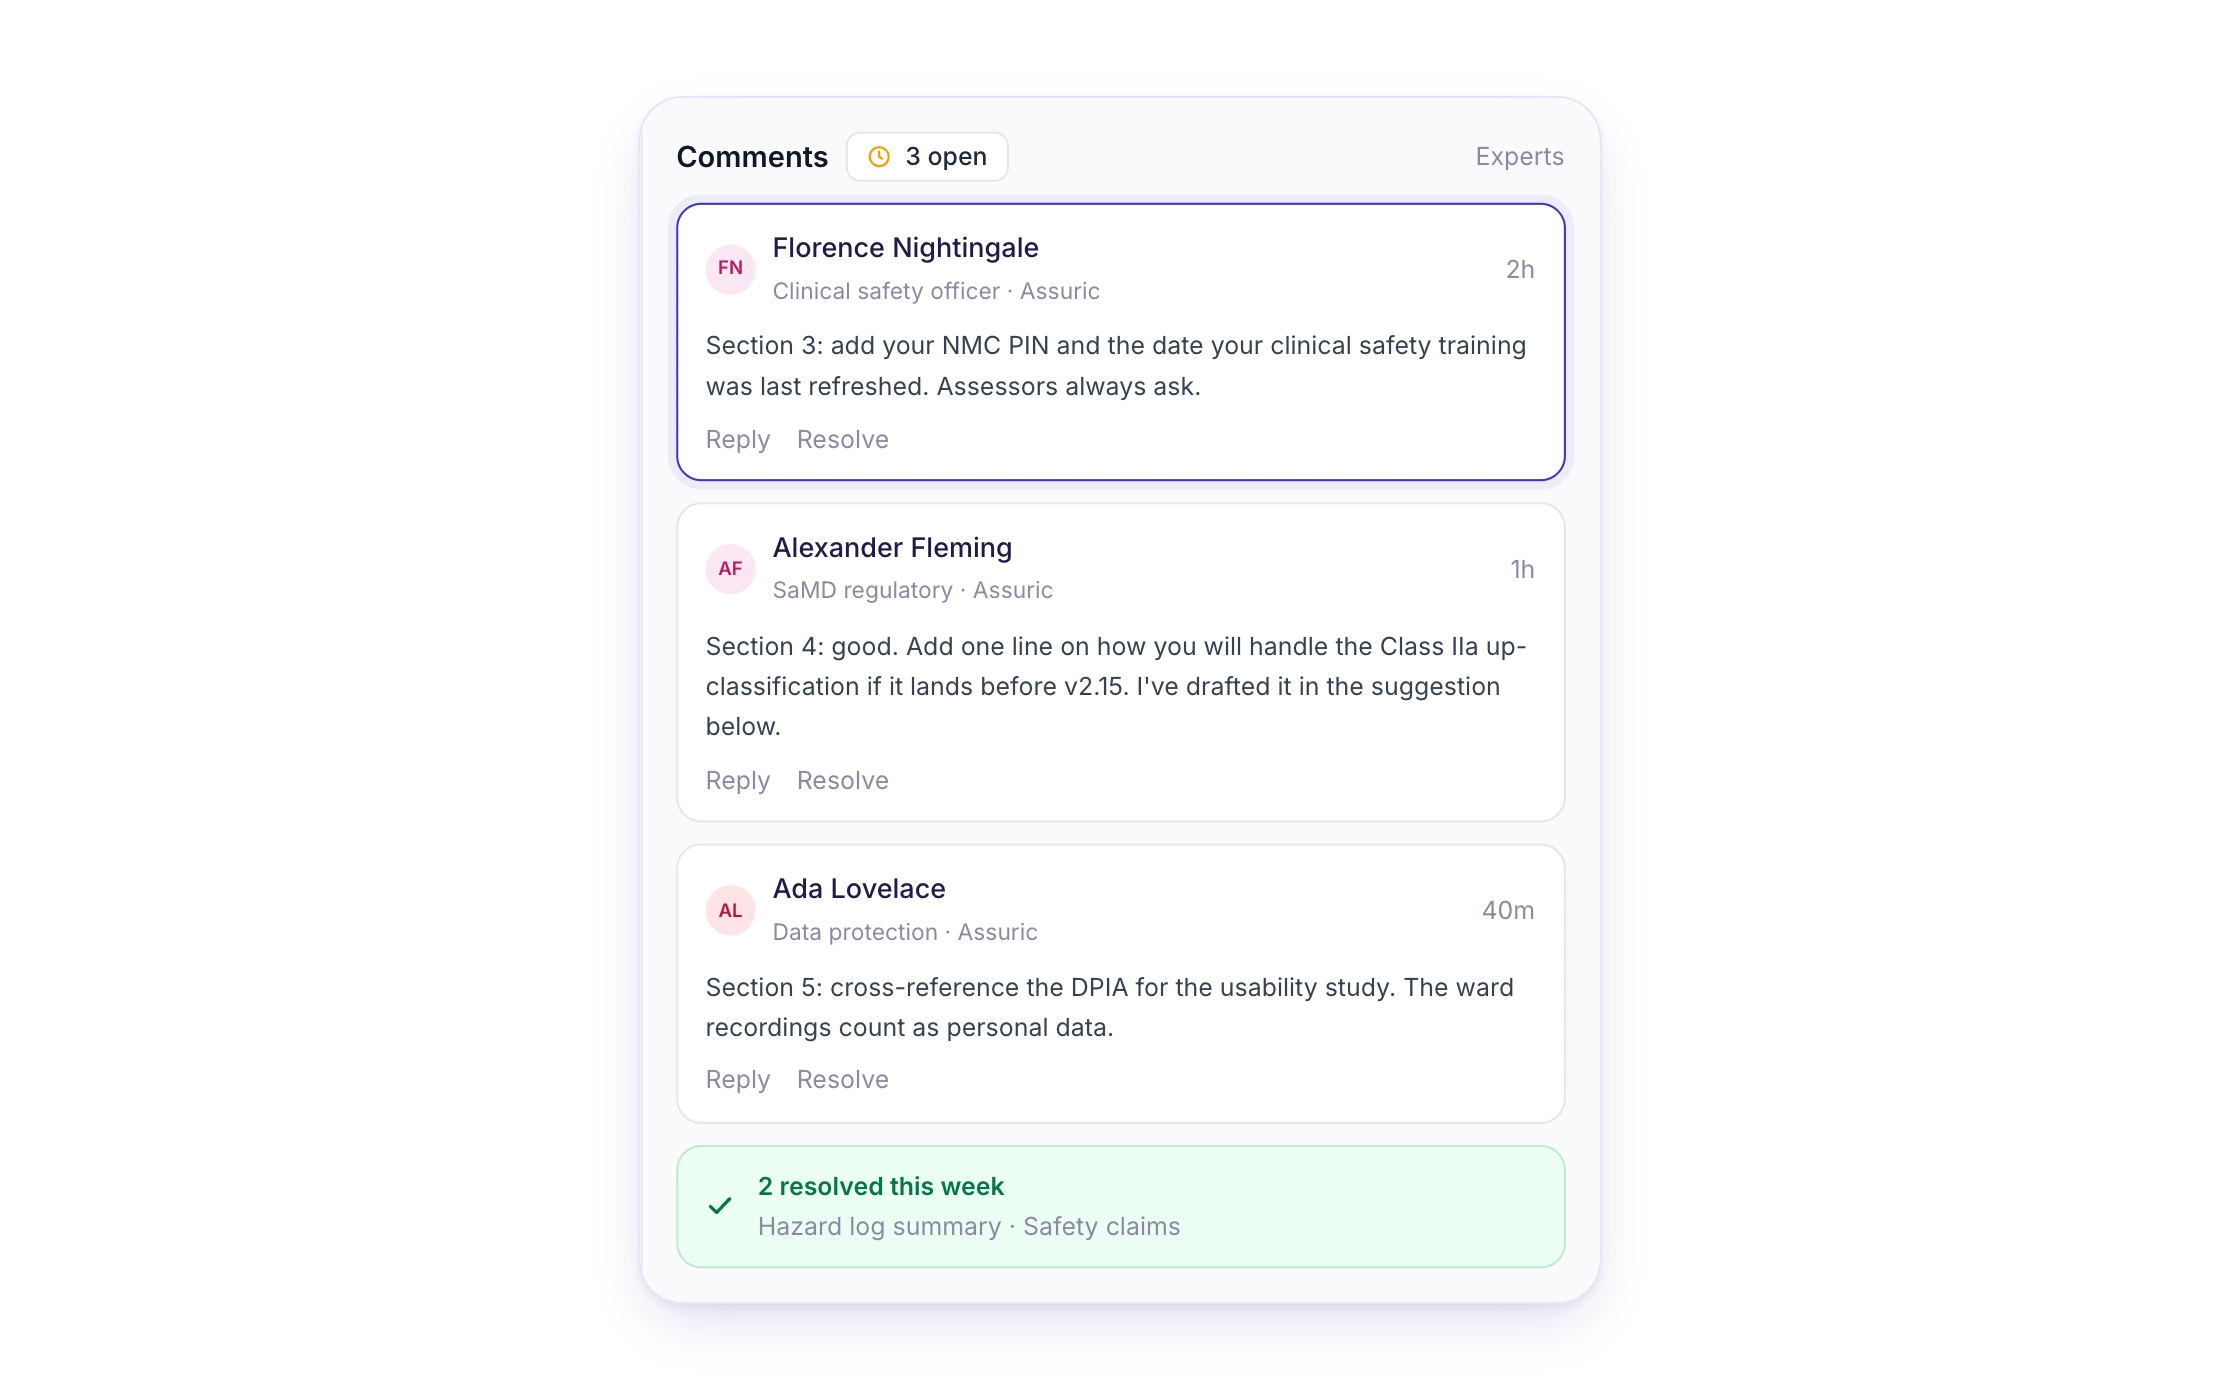
Expert hands-on support
Work with our regulatory team on your clinical evaluation plan, your literature and data appraisal and the report itself, then keep it current as the device changes.
Benefits
Assuric holds all three key requirements in one place, ensuring a record written once counts for ISO 13485, IEC 62304 and the MDR together rather than being rewritten across systems.
An eQMS that covers all SOPs, training records, CAPAs and internal audits comes pre-built to ISO 13485, with full controlled document management.
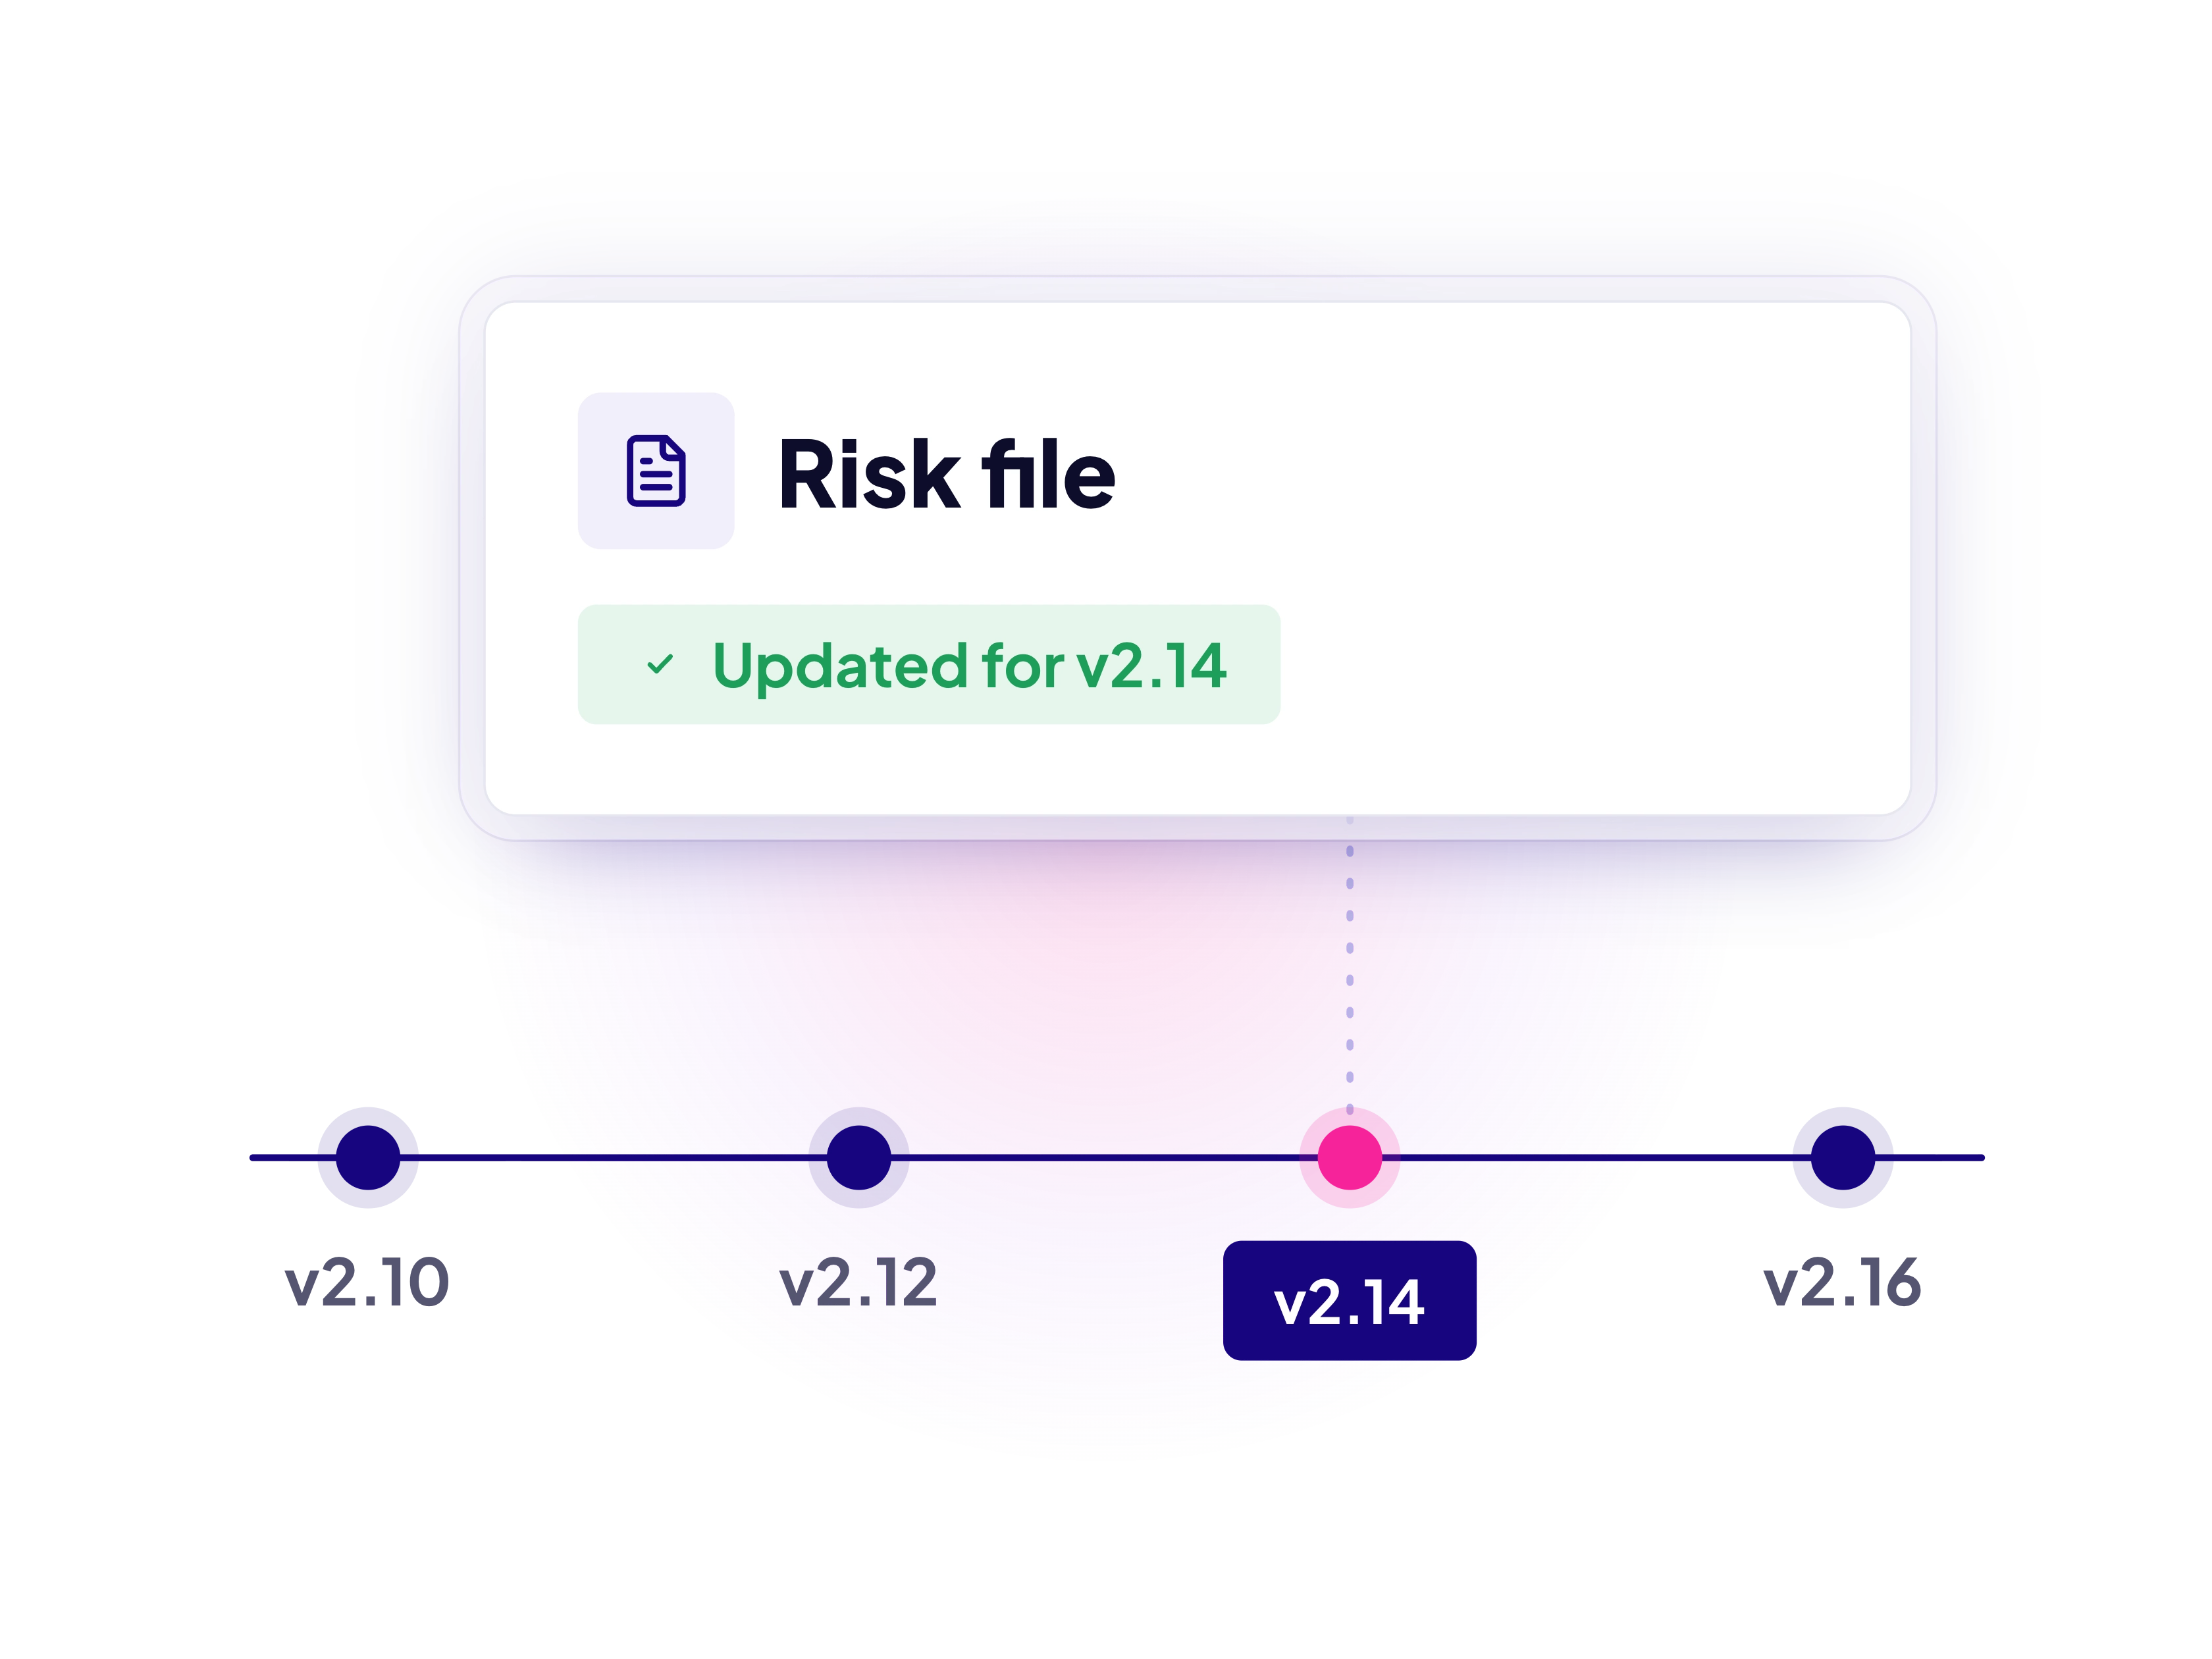
Ensure your risk file reflects the version you have shipped, with hazards and controls updated as the software changes and as field data comes in.
Frameworks
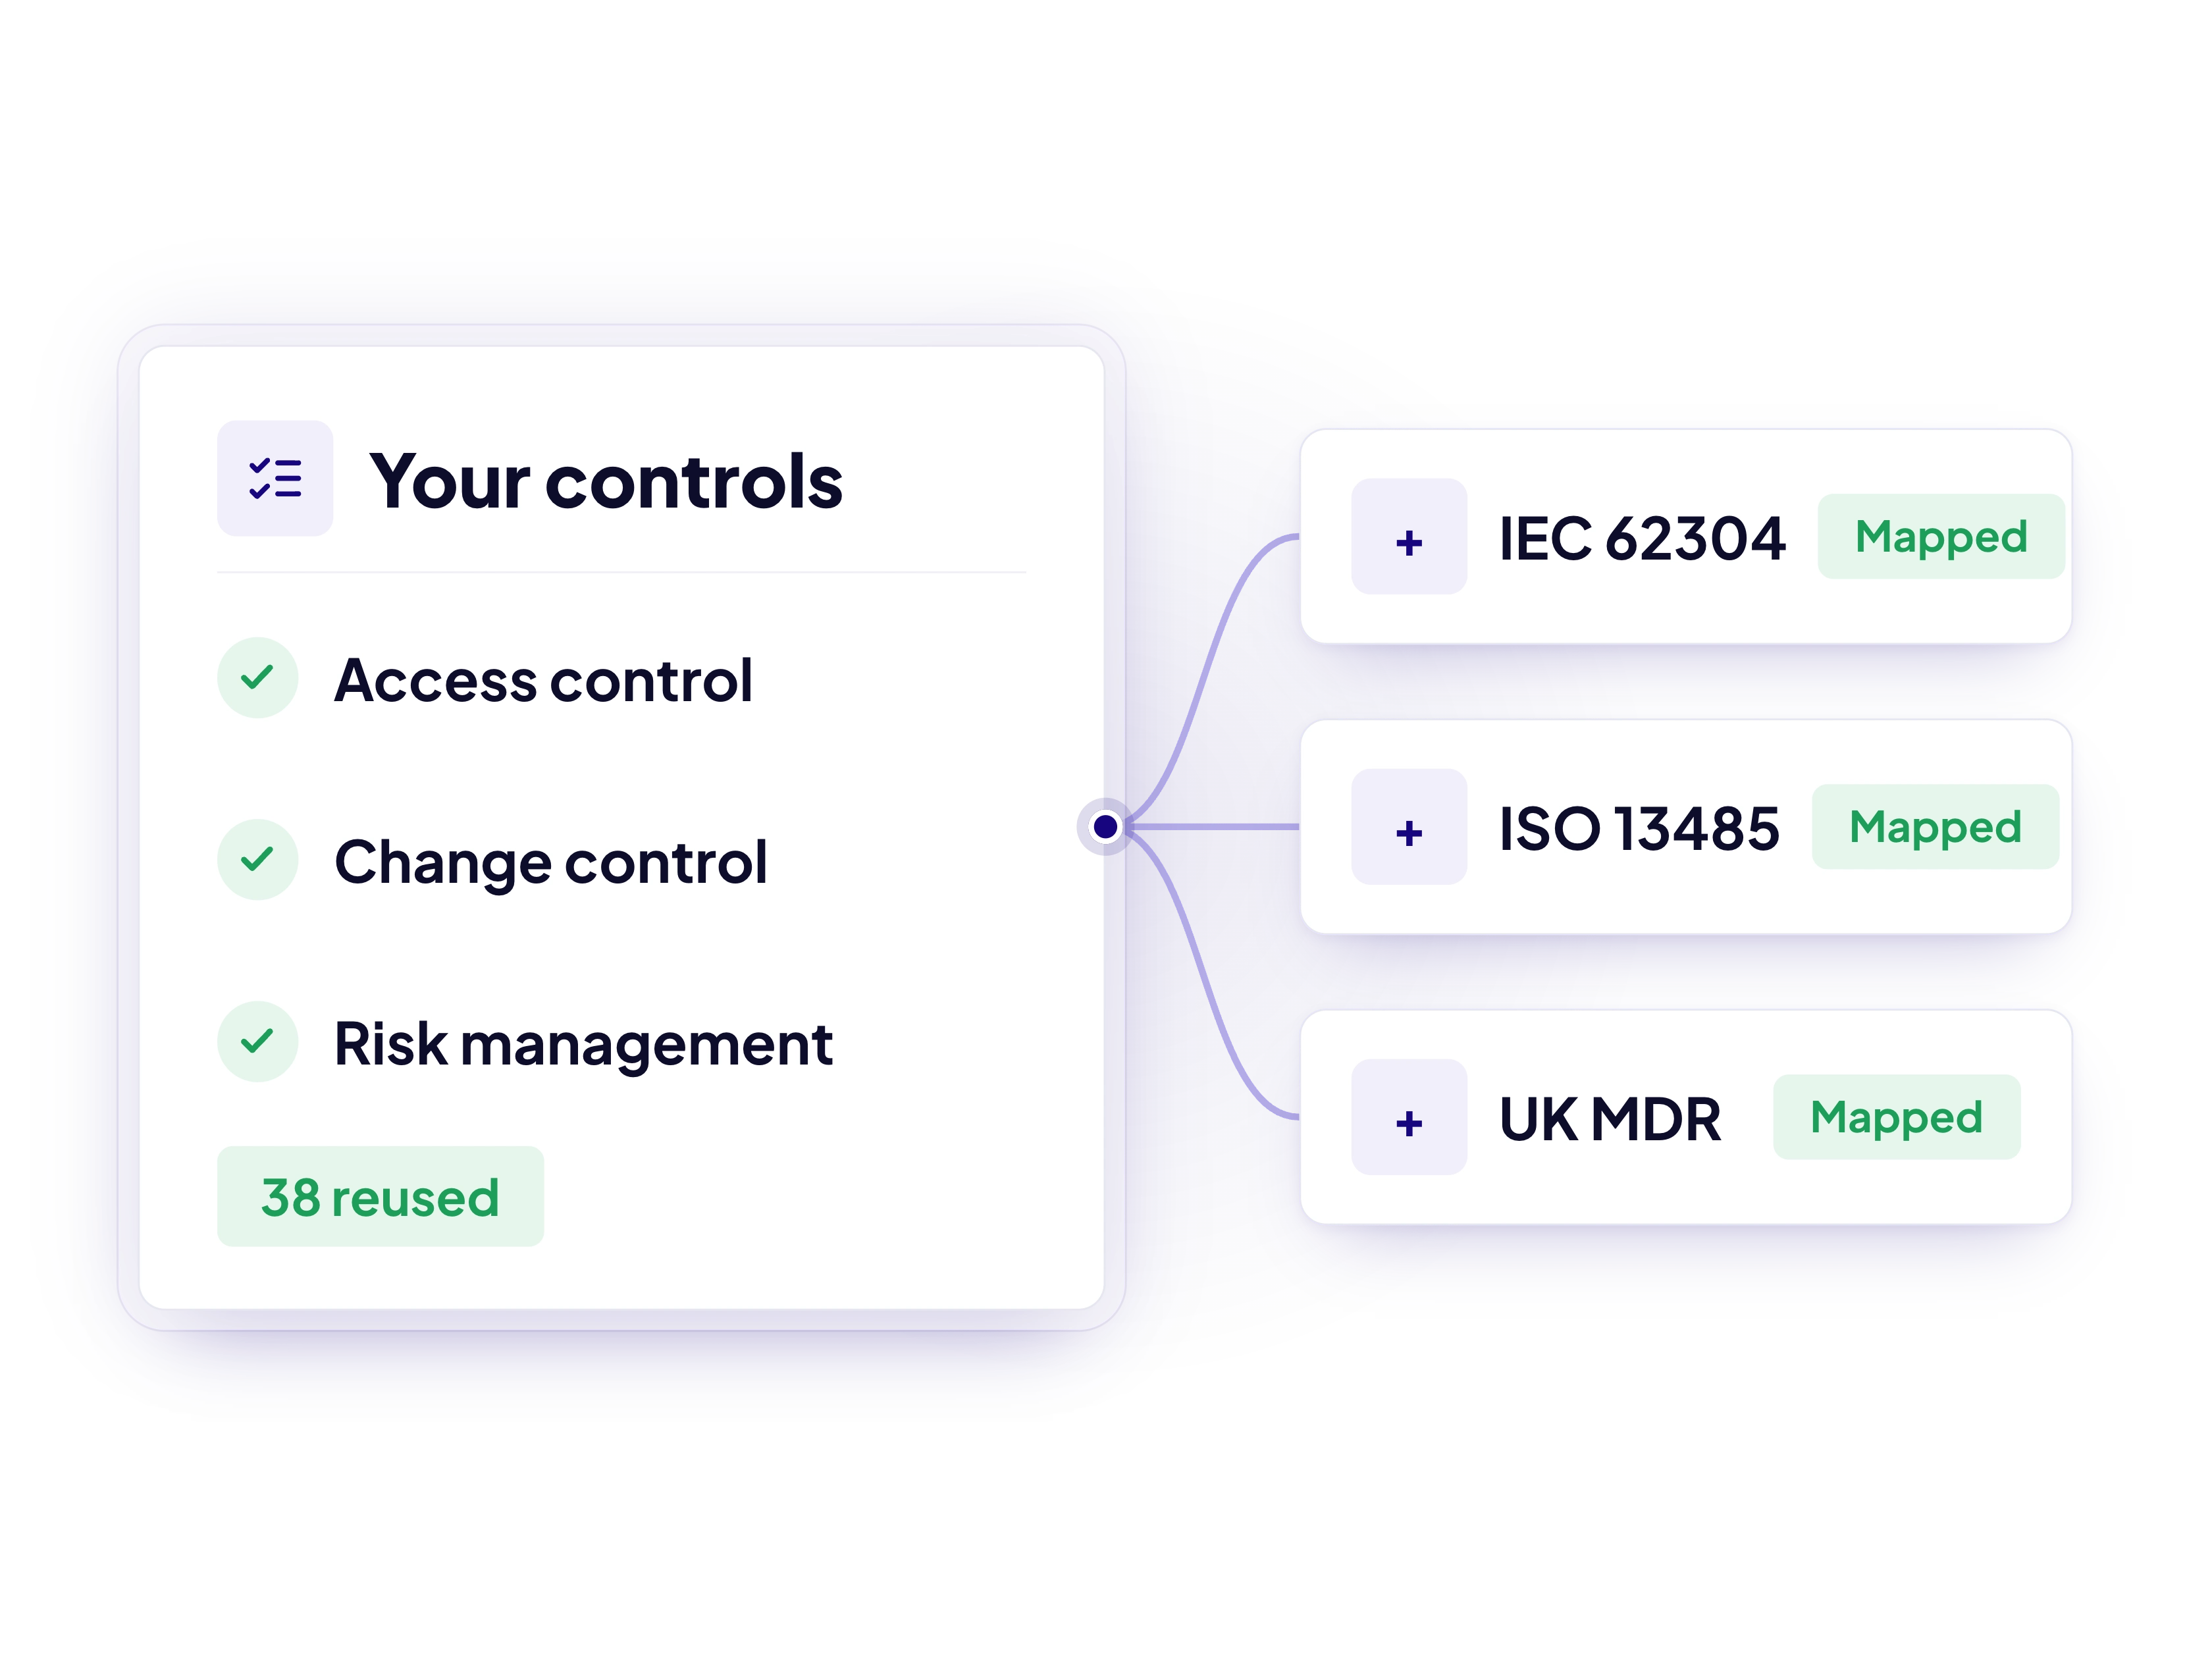
Overlapping requirements and evidence are mapped across frameworks automatically, so you never do the same work twice.

“We couldn't speak highly enough of both the Assuric team and the platform itself, which is incredibly easy to use, and with the skill and hardworking nature of the Assuric team. They've been a key component in accelerating our progress and deployments!”
Maks Kozarzewski
COO, VitVio

“The Assuric platform is a game changer in the clinical risk management field. It has been carefully designed to make the job of a CSO significantly more streamlined. The team are excellent and responsive to any product suggestions I have requested, and it has been a pleasure to work with them.”
Paul Llewellyn
CCIO, Accenture

“Assuric has been fantastic in helping us quickly and safely navigate regulatory compliance in the UK. From completing Cybersecurity requirements to DSPT, DCB0129, and DTAC, the team was supportive, extremely knowledgeable, and the platform made everything quick and straightforward. A separate regulatory company we consulted at the beginning even remarked on how quickly we achieved compliance!”
Katie Baker
Director UK & Ireland, Tandem
Regulation (EU) 2017/745, the Medical Device Regulation. It entered into force in May 2017 and has applied since 26 May 2021, replacing the Medical Devices Directive and the Active Implantable Medical Devices Directive. It sets out classification, conformity assessment, technical documentation, clinical evaluation, post-market surveillance and vigilance for any device placed on the EU market, and it applies to manufacturers outside the EU who sell into it.
It depends on your device class. Class I devices are generally self-declared by the manufacturer, while Class IIa, IIb and III devices need a notified body to assess your quality system and technical documentation before the CE mark goes on. Class I devices that are sterile, have a measuring function or are reusable surgical instruments need a notified body for those aspects.
The EU MDR governs the EU market and results in a CE mark. Great Britain is governed by the UK Medical Devices Regulations 2002 as amended, overseen by the MHRA, and results in a UKCA mark. The underlying expectations of a quality system, a risk file and technical documentation are close enough that the same evidence serves both, but they are separate routes and selling into both markets means meeting both.
Article 10 requires a quality management system, and ISO 13485 is the harmonised standard used to demonstrate one. It is not named as a legal requirement in its own right, but a notified body will expect a compliant QMS for anything above the lowest risk class, and building to ISO 13485 is the cleanest way to evidence it.
Annex I requires a risk management system established, documented and maintained across the whole lifecycle of the device, with risks reduced as far as possible and residual risk judged acceptable against the benefit. ISO 14971 is the standard used to demonstrate that, and Assuric holds the risk file alongside the quality system rather than in a separate tool.
Clinical evaluation is how you show, using clinical data, that your device is safe, performs as intended and has benefits that outweigh its risks. Every device under the EU MDR needs one. It is documented in a clinical evaluation report and kept up to date across the life of the device.
A post-market surveillance plan and system, a periodic safety update report or a post-market surveillance report depending on class, trend reporting, and vigilance reporting of serious incidents and field safety corrective actions to the competent authorities. The obligation applies across all classes, and the depth and frequency scale with classification.
Assuric holds your ISO 13485 quality system, your ISO 14971 risk file and the design records behind your technical documentation in one place. A record written once counts across ISO 13485, IEC 62304 and the MDR. Our regulatory team can also work with you on your clinical evaluation.
Your data is stored in the EU, in Belgium. Customer data is not used to train models.
Keep your QMS, risk management and technical documentation inputs in one place, ready for conformity assessment.


